Wednesday, October 12, 2011

Benzene Chromium Tricarbonyl and Challenge

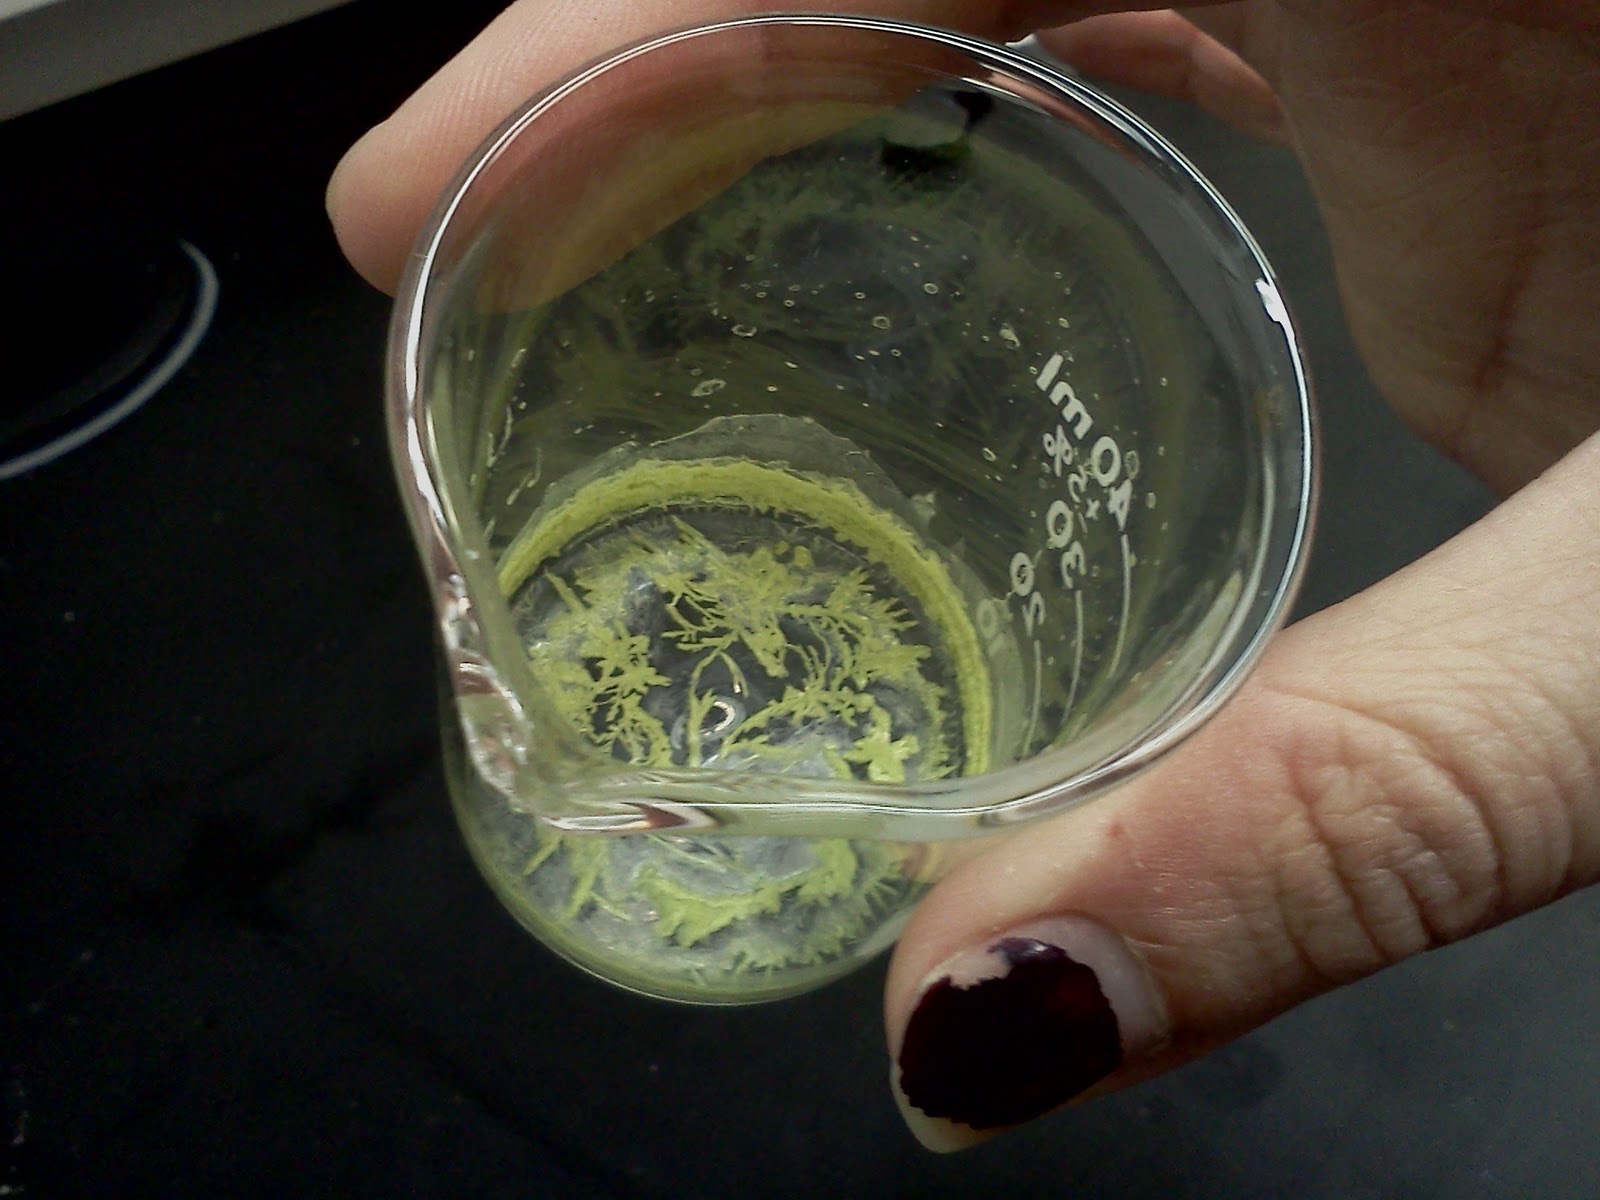
On Tuesday 10-4-11 we checked via IR to see if we had made our desired compound. First, we got an IR of the pure Cr(CO)6 in methylene chloride to compare to our experiment. There was only one carbonyl peak at 2000 1/cm. We pipetted out a small sample of our solution from the flask. There was both a solid yellow-green precipitate and a similarly colored liquid in the reaction flask. We used the liquid for the IR and got a spectrum that had two carbonyl peaks that were slightly lower than the peak on chromium hexacarbonyl. The values were very close to tabulated values for the complex, so we are confident that we got the desired complex. We then suction-filtered the contents of the flask to get a light green precipitate but there were also some black specks in the solution. We recrystallized our product by putting the precipitate in a mininum of methylene chloride and letting it sit overnight to evaporate the methylene chloride. The next day when we went to go check the product, we had green crystals in the beaker:


We also wanted to take a C-13 NMR and an H-NMR of our product. We did this by scraping out some of the crystals and placing them in an NMR tube with some deuterated DMSO. Our product dissolved in this and we got both NMR spectra. They both correspond very well to expected values for our complex.

We had to come up with a demonstration for National Chemistry Week that had to do with health, hygiene, or medicine. Purple Chrome decided to do a demonstration about acid erosion of teeth. Pepsi and other soft drinks contain phosphoric acid, so we wanted to test and see what prolonged exposure to your teeth actually does. We made a 2 M and a 7 M solution of phosphoric acid, a 2 M solution of NaF (which should theoretically make the tooth stronger, not weaken it), and we also had some Pepsi. We massed 4 teeth and placed one in each of the solutions. Immediately, the teeth in the acid and in the Pepsi started fizzing:

The tooth in the NaF didn't really do anything once it was put in the solution. We left the teeth sit in the solution for 4 and a half days, checking in on them periodically. We could see the erosion of the teeth in the acid happening very quickly. Even the tooth in the Pepsi wasn't safe from some acid erosion! We took them out on Tuesday 10-11-11 to dry, and then we massed them. The teeth that had been in the acid were very soft and almost disintegrated when we tried to get them out onto the watchglasses to mass them. Our hypothesis was correct that the teeth in the acid would lose mass due to erosion ("demineralization") and the tooth in NaF would gain some mass due to remineralization. Here is what each of them looked like after we took them out of the solutions:
Look at how the coloring in the Pepsi stained the tooth! It was just white before it sat in solution.

The tooth in NaF wasn't eroded like the teeth that were in the acid.

This tooth, before it sat in the acid, actually looked a lot like the tooth that was in NaF. See how the top of the tooth (facing right) is almost completely eaten away, and there are just deep craters remaining!

This one USED to be a whole tooth, but it was so soft and degraded that it fell apart when we tried to get it out of the vial of acid. In places, it was so thin it was translucent!


This experiment definitely made us think twice about drinking Pepsi!!


Wednesday, October 5, 2011

Not Quite Perfect

The next day, we went up to the lab after the solution had 20 hours to reflux to find that during the night, we had an air leak and all of our solvent had boiled away. Nothing was left except a black residue on the inside of the flask and all over the stir bar. We got this cleaned up and decided to start the reaction again on Thursday.

On Thursday 9-29-11 we set the reaction up in the same way except for two differences: we used a smaller flask so that the contents would have more volume in the bottom of the flask (hopefully this would prevent the solvent from boiling away too fast) and we used a two neck flask instead of a three neck flask.


This would have one less way for the solvent to escape, so the reaction has one less way to fail. We didn't want to leave the reaction to run overnight again in case it would fail, so we set it up at the end of lab, and one of us could come in on Friday morning to turn the heat on.

On Friday 9-30-11 we turned the heat on at about 10:30 AM and let it go all day until 4 PM. The heating mantle was turned to 3 this time instead of 4. We still got the black residue on the inside of the flask, but we could tell that we still had solvent in the flask because nothing had bubbled up through the joints and there was still liquid dripping from the condenser back into the flask.

On Monday 10-3-11 we started a second round of heating at 11 AM and didn't turn it off again until 5. The total cook time at this point was 11.5 hours, which is less than the 20 hours we originally decided to do but we decided to do an IR spectrum of the solution the next day during lab to see if the reaction had proceeded at all, or whether it had failed again.

Finishing Ferrocene and Beginning Benzene

FINALLY...we got our acetylation of ferrocene to work. We ran the GC on Tuesday 9-27-11 and the highest peak on the spectrum was monoacetylated ferrocene. That is the most compelling evidence we have, and the only evidence we have that still contradicts the presence of monoacetylated ferrocene is the low melting point we got. However, 3 out of 4 isn't bad! That successfully finishes the ferrocene lab, which is good because we have gotten tired of writing about it!

Along with finishing ferrocene, we finished our purple crystals lab on Tuesday. Allen was responsible for this while I (Emily!) was working on other things. He did the isolation like with the benzene inclusion compound, but this time it was just the pure [Ni(en)2(NCS)2] crystals without any benzene added to it. These crystals took so long to grow, and even after five weeks, there were still only a couple of crystals in the vial (nothing like the vial full of benzene crystals we got). We finally decided to purify them and take the melting point since they didn't seem to be growing any more crystals. After the isolation, we got some very pretty dark purple crystals:

These actually look nothing like the benzene inclusion crystals, which were not shiny and dark purple but more dull and light purple. They were also much smaller crystals than these. Here's a picture of the benzene crystals to jog your memory:

We massed the new crystals so we could find percent yield for the lab report, and then we took the melting point, which ended up being above 220 degrees Celsius (that is the limit on our thermometer, so we couldn't get more precise than that!).

Since those two things didn't take much time, we were able to set up our first reaction for our final project. We are doing our project on Cr-Arenes, in which Cr(CO)6 is reacted with an arene in a solvent mixture of dibutyl ether and THF. There are so many arenes to choose from, but we just picked 6 of them to try and synthesize, purify, and characterize. The first arene we wanted to try was benzene.

We set up a three neck flask exactly the same as in the synthesis of ferrocene except with a condenser on the middle neck of the flask:

The condenser is needed to air-cool the reaction so the solvent doesn't boil away. We vacuumed and argon-filled the flask three times and then left the argon running throughout the experiment. We then added in the two solvents via syringe. We added 8 ml of dibutyl ether and 1 ml of THF.


 Now we needed to bubble argon through the solvents to get rid of any oxygen that might ruin the reaction. We "adapted" a disposable syringe to fit into a hose from the rack and stuck the needle through the septum so that the argon bubbled through the solvents. It actually looked like the solvents were boiling because of the gas bubbles:


While this was bubbling, we got 1 g of Cr(CO)6 (which is a white solid) and measured out 1 ml of benzene in a syringe. We had to take the septum out before we could add these two reactants plus a stir bar, so we just replaced the septum with a greased glass stopper. We had to wire down all the attachments and the hoses to prevent any accidents. After that was done, we turned on the stirring mechanism on the hot plate, and turned the heating mantle on to 4. Once it started refluxing, the color of the solution changed quickly from clear with the white Cr(CO)6 floating around undissolved to a light yellow-green color:

Since we have been reading about these complexes, we have seen that they usually have a yellow or orangish color so this color change was a good sign! We left the lab on Tuesday night feeling pretty confident that we would just let it stir for 20 hours, and we would come back on Wednesday to a perfect product that we could purify and characterize.

Monday, September 26, 2011

Acetylation of Ferrocene, Part Two

On Thursday 9-22-11, we had to redo the acetylation part of the ferrocene lab because it did not work the first time. The GC of our product did not yield anything, so we decided to try a different procedure to make the same product. This new procedure was scaled up considerably so we hoped that there would be enough of our product to run tests on.

We began by adding ferrocene, acetic anhydride, and phosphoric acid in a round bottom flask and placing this flask in a sandbath with an air condenser hooked up to a steady flow of argon gas. We heated and stirred it until the temperature reached 100 degrees Celsius, and then heated it for 10 minutes after that.


When we took the flask off to empty the precipitate, we found that in addition to the precipitate, we had a tar-like black substance stuck on the bottom of the flask. It had actually formed around our stir bar, so the stir bar was probably not stirring for a long time before we realized it! We did have some precipitate so we washed that into a large beaker of ice and waited for the ice to melt before we added some sodium hydrogen carbonate until the solution stopped bubbling with the addition of the base. This took a huge amount of NaHCO3 (around 7 grams) and by this time, the solution was starting to look like we were just boiling a beaker of mud. Once in the ice bath, it started to look a little better. It looked like a brown precipitate in a reddish solution.


We suction filtered it and got a large amount of brown precipitate, which we then used to make a GC sample, an IR sample, and a Mel temp sample.


The trouble is, we didn't get the GC yet due to technical difficulties and the Mel temp gave a melting point of about 59 degrees Celsius while pure acetylated ferrocene has a recorded melting point of about 81 degrees Celsius.

We thought that maybe there were impurities in the sample causing the melting point aberration, so we decided to recrystallize our product. We did this by adding the minimum amount of petroleum ether to our sample to make most of it dissolve. We then decanted the clear yellowish liquid (which was the product dissolved in the pet ether) into a clean beaker and let it sit overnight. After the night was up, the ether had all evaporated off and a layer of orange crystals was left on the side of the beaker--this was our pure product.


Unfortunately, a Mel temp test of the pure product gave us the same results as the unpurified product, so at this point we need our GC to give us some proof of acetylation. We know from the drastically lower melting point that there is a new chemical formed from this reaction, we just don't know what it is!

Silicon Polymer (Bouncing Putty)

On Tuesday 9-20-11, we made a silicon polymer that looked like a clear gel and bounced like a bouncy ball. We started by combining diethyl ether and dimethyldichlorosilane in a round bottom flask. When we started adding some water dropwise into the mixture, the flask got very hot (even when we had it in an ice bath!) and emitted some fumes. The fumes were actually hydrogen chloride gas, which is given off in the reaction we were doing-- hydrolysis of dimethyldichlorosilane. The product of the hydrolyzation (a silanol) was what we needed to make our putty. Of course, now there were two layers in the flask so we had to hearken back to organic chemistry and grab a separatory funnel.


We wanted to keep the organic layer, so we separated, washed the aqueous layer with hexanes, separated again, and combined the two organic phases. We used sodium hydrogen carbonate to neutralize the HCl that was formed in the reaction.

We dried the product over magnesium sulfate and filtered it. Then, we got to use the rotary evaporator to evaporate off the ether and obtain our product. Here are a few pictures of the roto vap:


We put our sample in the roto vap, filled the condenser up with ice, and turned it on. It heated the sample and evaporated the ether from it. The ether then went over to the condenser, where it was turned from gas to liquid and was collected in the round flask at the side. This left just our product in the vial inside the roto vap.


After this, we took the IR of the oily product and massed it to find out how much boric acid to add to the liquid (we needed about 7% by mass). When we added the boric acid, it just sat on the bottom of the beaker and wouldn't dissolve into solution. When we heated it with continual stirring, it started to get thick and gummy after about 15 minutes.

Once it seemed like all the liquid had turned to putty, we scraped it out of the beaker, put it on a preweighed watchglass, and found the mass. If it was left to sit too long on the watchglass, it would "melt" into a liquidy puddle but then would form right back into a putty again if you rolled it in your hands. Here is our bouncing putty...which actually did bounce!!

Friday, September 16, 2011

More Ferrocene and A Bit of Spectroscopy

The first thing we did on Thursday 9-15-11 was look at the product of our acetylation reaction that had been sitting for two days. Surprisingly, the solution was no longer blue-green, but had turned completely orange and there appeared to be an orange precipitate on the bottom (along with what looked like some excess NaHCO3).


We vacuum-filtered this and below is the picture of the orange precipitate. We then let it dry in the oven for a little bit, and it turned a light brown after this.

The yield was very small, but then the amounts of reactants were small as well. We barely had enough for a sample to be dissolved in methylene chloride to run through the GC-MS. We can only know if the acetylation of ferrocene worked once we get the spectra back from the GC-MS!

The purification of our ferrocene sample by sublimation was a really neat thing to watch. We put our crude sample in a glass petri dish on top of a hot plate and turned the hot plate on. We placed a beaker full of ice on top of the petri dish.


 As the sample heated up, we began to see little bright orange crystals forming on the inside of the petri dish lid under where the ice beaker was placed.


Finally, after about 20 minutes, there were crystals all over the sides of the petri dish bottom and all over the top of the petri dish. What was left of the crude sample was a black pile of dust:

Here is a picture of the top of the petri dish, which is definitely nicer to look at than the bottom:
We had to scrape all the crystals out of the petri dish and mass them so we could calculate percent yield.

The rest of the lab period involved getting spectra to see if we actually made what we thought we made. We ran the ferrocene crystals through the Mel-temp device and got a melting point very close to that of pure ferrocene, so it's looking good so far! We also ran an IR, H-NMR, and a GC-MS on ferrocene. We left the GC-MS to run overnight, so we don't have those spectra yet but we do have the IR and NMR:
IR Spectrum of our Product

The only thing that was weird about this was the carbonyl peak that shows up at about 1750 1/cm. We ran background scans several times, but it never showed up in the background scan, just in the spectrum of our product.

H-NMR Spectrum of our Product:
The single peak at about 4 ppm is characteristic of ferrocene because of the aromatic cyclopentene rings on either side of the Fe atom in the "sandwich compound":
                        

Structure from http://chemistry.about.com/od/factsstructures/ig/Chemical-Structures---F/Ferrocene.htm

So far, we seem to have a lot of evidence in  favor of us actually making ferrocene, so we'll see if the GC-MS disputes that or supports it.

Finally...Ferrocene!

This Tuesday (9-13-11) we tried the ferrocene lab again, but this time we used a different procedure. The reaction stayed the same except for the solvent and the amount of reactants. In the last one, we used ethylene glycol but in the new procedure we used 1,2-dimethoxyethane (also known as "Glyme") as the solvent. This reaction was scaled up considerably--in the last one, we used 2 g of KOH and in this one we had to use 7 g of KOH (which is a lot more than is pictured below!). We were worried that the large amount of KOH might not dissolve and the reaction would fail, but it seemed to dissolve quickly.


The whole setup of the reaction went the same as the last one, so we had some practice and the step where we put the flask under a vacuum and inert argon atmosphere went a lot faster this time.


We didn't really notice anything changing from the last reaction to this reaction until we added the FeCl2/DMSO solution using the cannula. The last time, it was a very pretty dark red color but this time it was more of a dark brown color. The biggest change we noticed is when the HCl and ice was added at the end. The last time, the solution immediately turned a bright lime green (and was almost completely clear) but this time, it turned a murky dark blue-green color. The reason it was murky? The bunch of orange solid that precipitated out, AKA ferrocene!!!! At least, we really hoped that our spectroscopic evidence would tell us that it was actually ferrocene. What the heck, we were just excited that the reaction had produced SOMETHING! We vacuum filtered the solid, dried it in the oven, and massed the chunky reddish-brown solid that it had become.

That same day, we tried the acetylation of ferrocene reaction with commercial ferrocene that was available in the lab (we were trying to save time since we lost a day with the first ferrocene experiment). The only reactants that went into this reaction were ferrocene, phosphoric acid, and acetic anhydride. This stirred at 100 degrees celsius for 10 minutes under an air cooled condenser. The mixture changed from a light orange to a dark brown color at the end of the heating. After this, NaHCO3 was added in small increments, the key word here being "small" because each time we added some, the mixture bubbled very high in the flask and expelled CO2 gas. After the mixture stopped bubbling, it had turned a strange blue-green color but we didn't see any evidence of the brown solid we were supposed to be making. We set it in an ice bath, and then let it sit at room temperature until Thursday since we were out of time. We weren't feeling too confident about this reaction, but we would have to wait and see what happened with it!